
GROMACS的PME网格与截断参数实操
216个SPC/E水的实际GPU短MD:完整输入、终端操作、数据与图。
阅读本篇目录
下载16组完整输入、原始日志、能量数据与绘图脚本。所有例子使用同一盒216个SPC/E水分子(648原子),在鸢算科技集群P100上实际运行20 ps。GROMACS 2026.4使用CUDA 12.8、thread-MPI构建;每个任务使用1张GPU、2个CPU线程。建模与最小化见Packmol接GROMACS水盒工作流。
运行步骤#
先解压下载包,进入gromacs-water子目录。下面各个算例互相独立,每次从下载包根目录进入对应目录;为每个需要运行的目录保存一份下方的run.sh。start.gro包含坐标和速度,topol.top引用安装包自带SPC/E参数。下面的脚本在所选算例目录保存为run.sh:
#!/bin/bash
#SBATCH --partition=gpu-8p100-hs
#SBATCH --nodes=1
#SBATCH --ntasks=1
#SBATCH --cpus-per-task=2
#SBATCH --gres=gpu:1
#SBATCH --time=00:20:00
set -eo pipefail
source /etc/profile
module purge
module load gromacs/2026.4-p100-threadmpi
export OMP_NUM_THREADS=2
gmx grompp -f md.mdp -c start.gro -p topol.top -o md.tpr
gmx mdrun -deffnm md -ntmpi 1 -ntomp 2 -nb gpu -pme gpu -bonded cpu -update cpu
if grep -Eq '^pcoupl[[:space:]]*=[[:space:]]*no([[:space:]]|$)' md.mdp; then
  printf 'Potential\nTotal-Energy\nTemperature\nPressure\n0\n' | gmx energy -f md.edr -o thermo.xvg
else
  printf 'Potential\nTotal-Energy\nTemperature\nPressure\nVolume\n0\n' | gmx energy -f md.edr -o thermo.xvg
fi
sbatch run.sh
# 任务结束后检查最后几行日志
 tail -n 20 md.log
constraints = none不代表这里的水是柔性水:SPC/E拓扑含SETTLE,水分子内部几何仍保持刚性。continuation = yes与gen-vel = no保留输入速度。本组从同一GRO起点分叉,GRO速度有输出精度限制,并不是读取同一完整检查点的位级重启。
本例比较什么#
基准fourierspacing为0.12 nm、rcoulomb和rvdw均为0.8 nm。分别修改网格间距或两个截断,其他输入保持同一套设置,观察能量和压力变化。
nvt-v-rescale#
cd nvt-v-rescale
sbatch run.sh
完整 md.mdp:
integrator = md
dt = 0.001
nsteps = 20000
continuation = yes
cutoff-scheme = Verlet
coulombtype = PME
rcoulomb = 0.8
rvdw = 0.8
pbc = xyz
tcoupl = v-rescale
tc-grps = System
tau-t = 0.1
ref-t = 300
pcoupl = no
gen-vel = no
constraints = none
nstenergy = 10
nstxout-compressed = 100
fourierspacing = 0.12
pme-spacing-0.1#
cd pme-spacing-0.1
sbatch run.sh
完整 md.mdp:
integrator = md
dt = 0.001
nsteps = 20000
continuation = yes
cutoff-scheme = Verlet
coulombtype = PME
rcoulomb = 0.8
rvdw = 0.8
pbc = xyz
tcoupl = v-rescale
tc-grps = System
tau-t = 0.1
ref-t = 300
pcoupl = no
gen-vel = no
constraints = none
nstenergy = 10
nstxout-compressed = 100
fourierspacing = 0.1
pme-spacing-0.16#
cd pme-spacing-0.16
sbatch run.sh
完整 md.mdp:
integrator = md
dt = 0.001
nsteps = 20000
continuation = yes
cutoff-scheme = Verlet
coulombtype = PME
rcoulomb = 0.8
rvdw = 0.8
pbc = xyz
tcoupl = v-rescale
tc-grps = System
tau-t = 0.1
ref-t = 300
pcoupl = no
gen-vel = no
constraints = none
nstenergy = 10
nstxout-compressed = 100
fourierspacing = 0.16
cutoff-0.65#
cd cutoff-0.65
sbatch run.sh
完整 md.mdp:
integrator = md
dt = 0.001
nsteps = 20000
continuation = yes
cutoff-scheme = Verlet
coulombtype = PME
rcoulomb = 0.65
rvdw = 0.65
pbc = xyz
tcoupl = v-rescale
tc-grps = System
tau-t = 0.1
ref-t = 300
pcoupl = no
gen-vel = no
constraints = none
nstenergy = 10
nstxout-compressed = 100
fourierspacing = 0.12
cutoff-0.7#
cd cutoff-0.7
sbatch run.sh
完整 md.mdp:
integrator = md
dt = 0.001
nsteps = 20000
continuation = yes
cutoff-scheme = Verlet
coulombtype = PME
rcoulomb = 0.7
rvdw = 0.7
pbc = xyz
tcoupl = v-rescale
tc-grps = System
tau-t = 0.1
ref-t = 300
pcoupl = no
gen-vel = no
constraints = none
nstenergy = 10
nstxout-compressed = 100
fourierspacing = 0.12
实际结果#
| 条件 | 10–20 ps平均温度/K | 温度时间波动标准差/K | 平均压力/bar | 最终体积/nm³ |
|---|---|---|---|---|
| nvt-v-rescale | 300.831 | 12.266 | 356.27 | 6.461670 |
| pme-spacing-0.1 | 300.536 | 11.649 | 610.08 | 6.461670 |
| pme-spacing-0.16 | 299.507 | 12.194 | 597.03 | 6.461670 |
| cutoff-0.65 | 300.489 | 11.824 | 1102.55 | 6.461670 |
| cutoff-0.7 | 300.231 | 11.833 | 551.27 | 6.461670 |
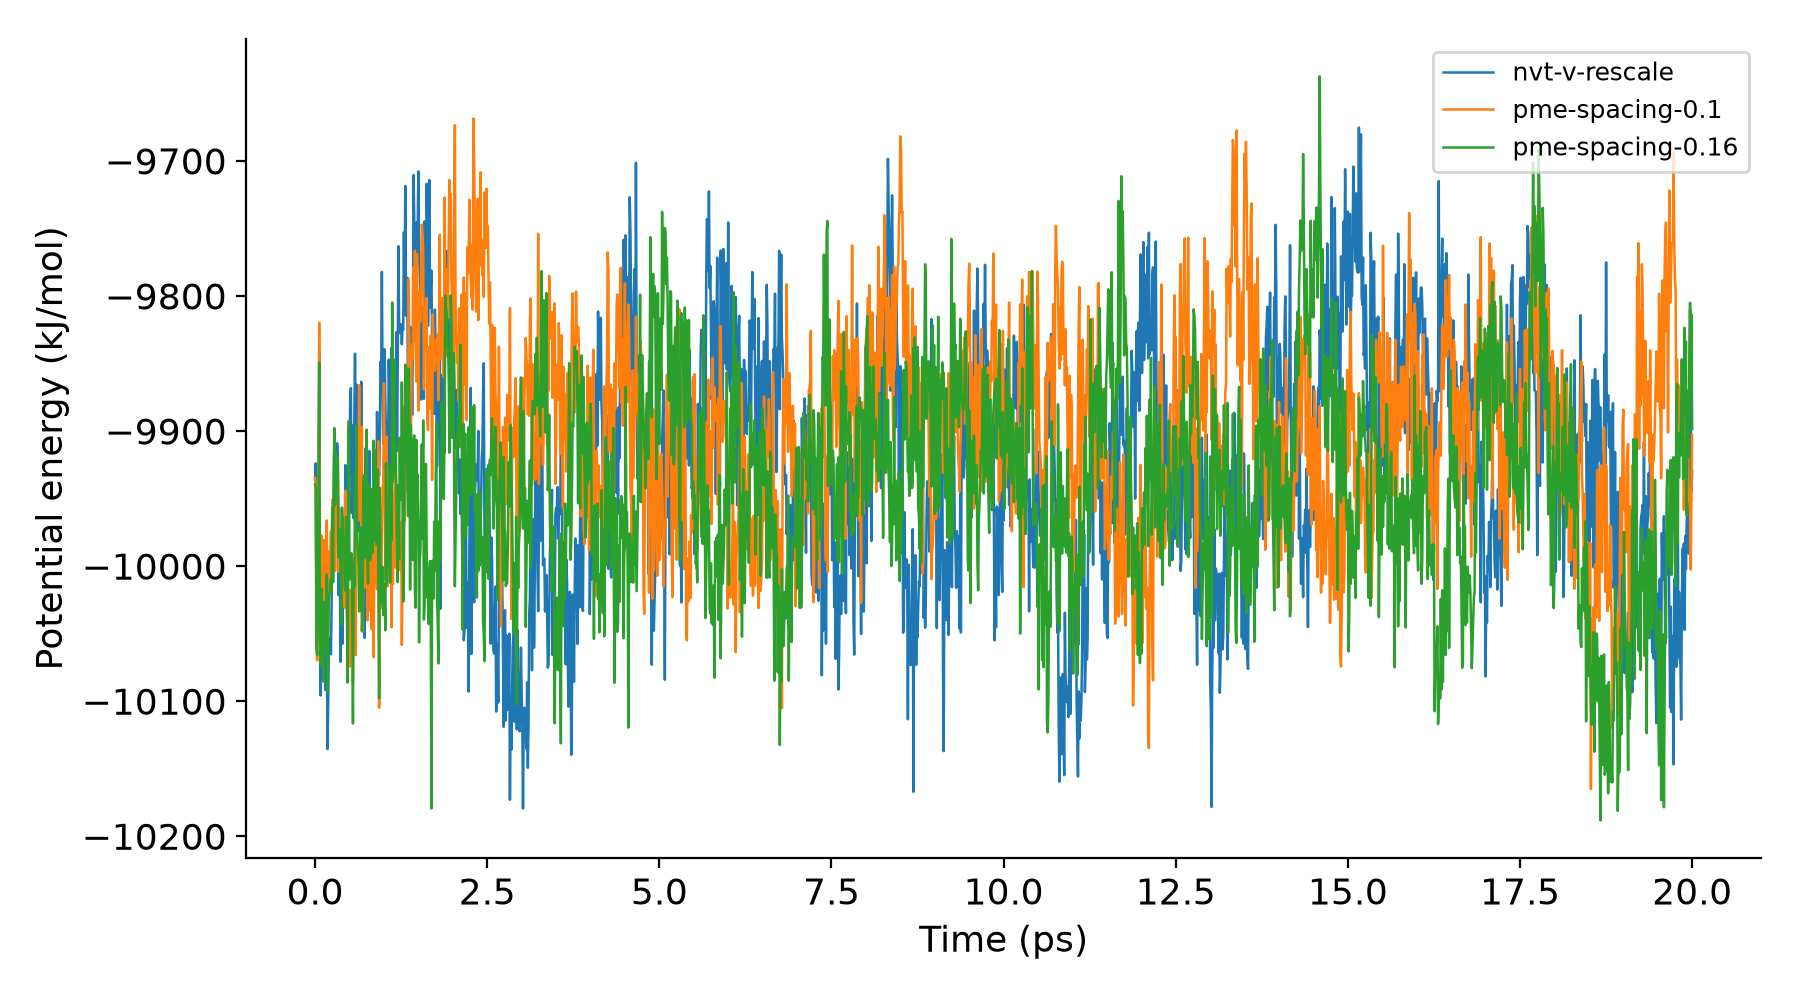
fourierspacing是目标网格间距,实际FFT网格按盒长与实现要求取整,实际设置应看md.log。不同参数的轨迹会逐渐分离,本例是操作示范,不是同一批冻结构型上的纯数值误差分解,也不是性能排名。生产参数应按所用力场要求选择并检查收敛,不能仅选最快的一组。
表中的标准差是轨迹自身的时间波动,不是平均值误差条。
提取与画图#
返回下载包根目录:
module purge
module load gpumdkit/1.5.8
python analyze.py
得到analysis/RESULTS.json和六幅PNG曲线。脚本按XVG列名读取数据;定容任务的体积从GRO盒矢量取得。NPT若重新提取体积,使用:
printf 'Volume\n0\n' | gmx energy -f md.edr -o volume.xvg
这些例子用于学会修改输入、提交任务、检查输出和解读曲线。换成蛋白、界面或其他力场时,需要重新选择合理的截断、耦合时间与模拟长度。